
What Is Dissolution?
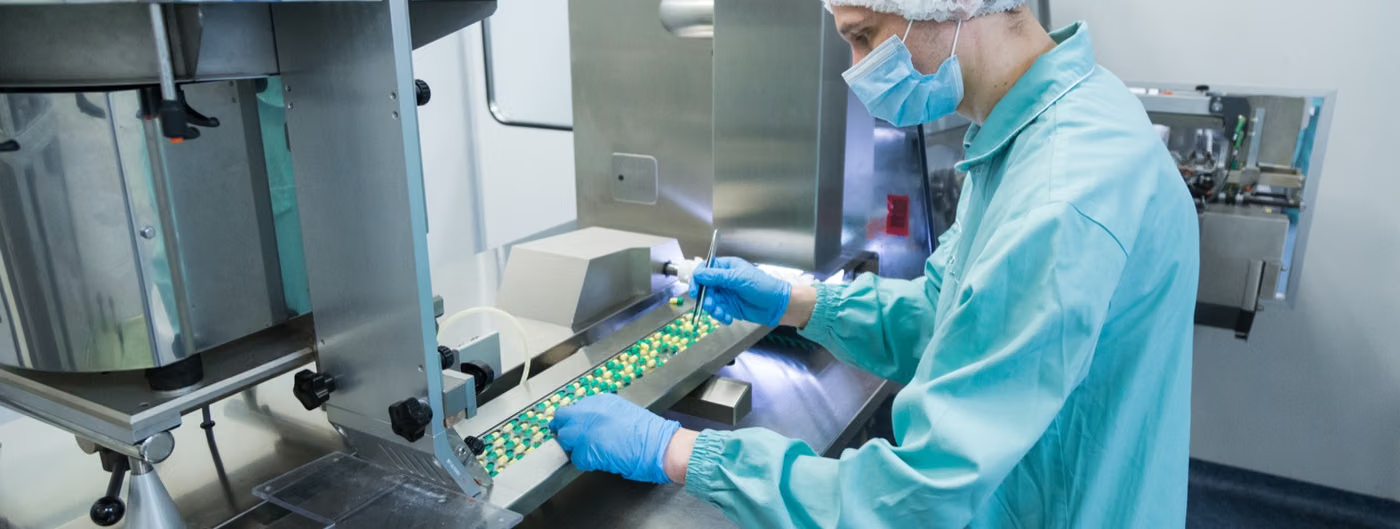
The dissolution test measures how much active substance dissolves from a tablet or capsule in a defined medium over a defined time. The specimen is placed in the dissolution medium at constant temperature and stirring speed; samples are pulled at defined time points and the active substance amount is measured.
Purpose of Dissolution
The purpose is to verify the product's active-substance release behaviour. The dissolution profile is an important indicator of bioavailability and is used to compare generic products against the reference product.
Which Products Is Dissolution Applied To?
- Tablets and capsules
- Extended-release forms
- Enteric-coated products
- Suspensions
- Transdermal systems
In Which Sectors Is Dissolution Used?
Standards Related to Dissolution
- Ph. Eur. 2.9.3 (dissolution test for solid dosage forms)
- Ph. Eur. 2.9.4 (dissolution test for transdermal patches)
- USP <711> (dissolution)
- USP <724> (drug release)
- ICH Q6A (specifications)
Dissolution Conditions
- Apparatus
- Basket (Apparatus 1), paddle (Apparatus 2), flow-through cell and others; set by the monograph
- Environment
- Dissolution medium, volume and pH defined in the monograph
- Temperature
- Temperature representing body temperature, as defined in the standard
- Stirring speed
- Rotation speed defined in the monograph
Equipment for Dissolution
- Dissolution tester and apparatus sets
- Automatic sampling system
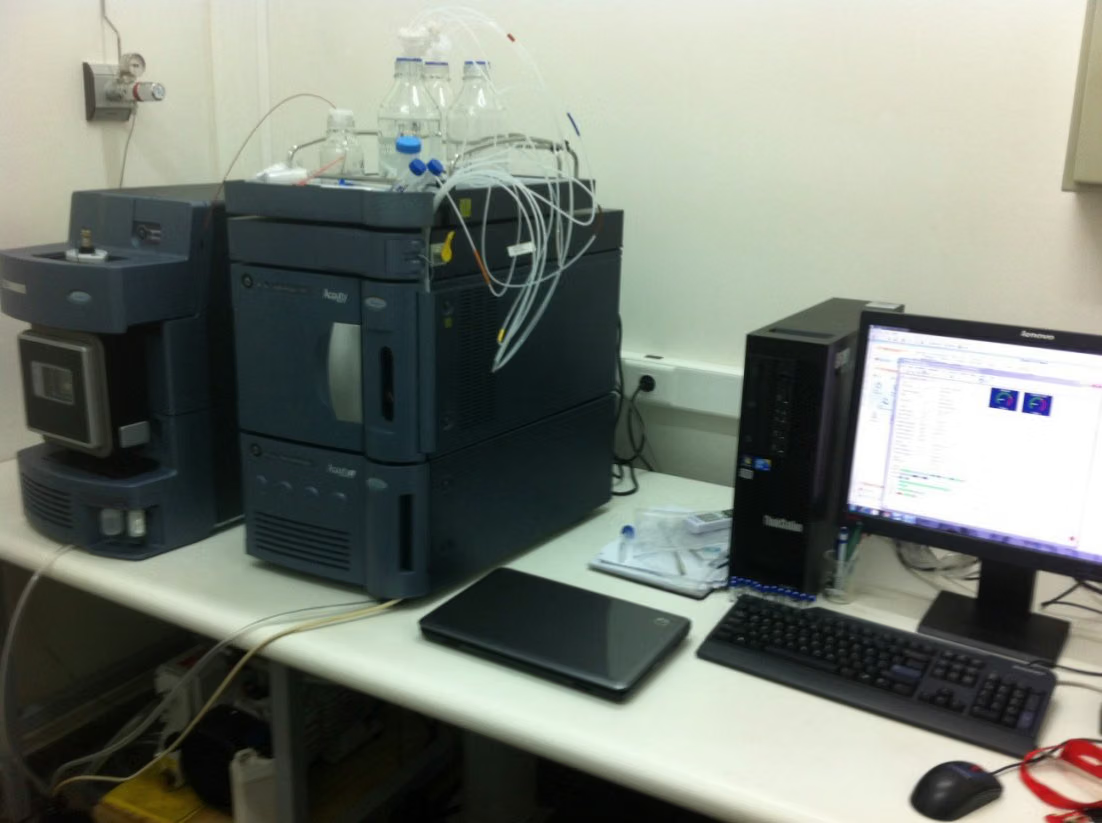
- HPLC and UV/Vis spectrophotometer
- pH meter and degassing equipment
- Calibration and mechanical qualification apparatus
Sample Requirements for Dissolution
- The number of units specified in the monograph (usually 6 or 12)
- Batch number and declared dose must be provided
- The reference product must also be supplied for a comparative study
- The specimen must be sent in its original packaging
Duration of Dissolution
The test is short for immediate-release products; profiling extended-release forms takes hours. Comparative profile studies take longer since they are repeated in multiple pH media.
Dissolution Procedure
- 1The apparatus's mechanical qualification and calibration are verified.
- 2The dissolution medium is prepared, degassed and brought to temperature.
- 3Specimens are placed into the vessels.
- 4The test is started at the defined stirring speed.
- 5Samples are pulled at the defined time points.
- 6The active substance amount is measured and the percentage dissolved is calculated.
Dissolution Acceptance Criteria
The acceptance criterion is not given on this page: for every active substance and every product, the limit comes from the relevant pharmacopoeia monograph or the approved specification in the marketing authorisation dossier. The pharmacopoeia's general chapter defines the method; the monograph sets the limit. The pharmacopoeia defines a staged acceptance approach (S1, S2, S3) for assessing dissolution results: if the criterion is not met at the first stage, the test is extended with additional units.
Evaluation of Results
The apparatus's mechanical qualification is the largest source of variability in the result: paddle wobble, vibration or dissolved gas in the medium markedly affect it. In comparative profile studies, similarity is assessed with defined statistical measures.
Dissolution Report
- Product identification, batch and declared dose
- Apparatus, medium, volume, temperature and stirring speed
- Percentage dissolved for each unit and each time point
- Mean profile and spread
- Acceptance stage (S1/S2/S3) assessment
Accreditation
EUROLAB holds internationally recognised accreditation obtained from accreditation bodies party to the IAF and ILAC multilateral recognition arrangements, under ISO/IEC 17025 for laboratory testing, ISO/IEC 17020 for inspection and surveillance, and ISO/IEC 17021-1 and ISO/IEC 17065 for system and product certification. This chain ensures the reports and certificates issued are accepted worldwide.
Whether this test can be offered within accredited scope depends on your product and the standard applied; since accreditation is method-based, the scope is confirmed in writing at the quotation stage.
Scope and authorisationsRelated Regulations
- 2001/83/EC Directive on Medicinal Products for Human Use
- GMP Guide (EudraLex Volume 4)
Pharmaceutical Analysis — What Does This Testing Field Cover?
The section below describes the field that the Dissolution test belongs to; the conditions and acceptance criteria of this particular test are given in the sections above. The scope specific to your product is confirmed in writing at the quotation stage.
Pharmaceutical analysis determines the identity, purity, quantity and behaviour of an active substance and of the finished product. What sets this family apart is that the methods are largely defined in PHARMACOPOEIAS: the European Pharmacopoeia (Ph. Eur.) and the United States Pharmacopeia (USP) give both the method and the acceptance framework together.
The binding nature of the pharmacopoeial method shapes how the field works. In other sectors a laboratory can choose a suitable method, whereas here the method in the monograph governs; if an alternative method is to be used, its equivalence must be demonstrated. This is what makes results comparable worldwide.
Analyses are carried out in several groups. Identity and assay (HPLC, UV, titration) show what the product is and how much active substance it contains. The impurity profile and residual solvents look for unwanted compounds arising from the manufacturing process. Elemental impurities address metals originating from catalysts and equipment. Dissolution and disintegration behaviour is the most important in vitro indicator of how the drug will be released in the body.
The DISSOLUTION TEST is one of the most critical measurements in the field. It is not enough for a tablet to contain the right amount of active substance; that substance must dissolve at a suitable rate. The dissolution profile is used to assess the effect of formulation changes on bioavailability and holds a central place in generic drug licensing.
METHOD VALIDATION is a mandatory step in this family. If a pharmacopoeial method is used, it is verified that the method works properly under that laboratory's conditions. If an in-house method has been developed, it is validated against parameters such as linearity, precision, accuracy, specificity, limits of detection and quantification, and robustness. A result produced with an unvalidated method cannot be used in a licence dossier.
Stability studies determine shelf life and are the longest-running work in the field. The product is held under defined temperature and humidity conditions, analysed at set intervals, and the results become the basis of the shelf life and storage condition claims. Accelerated conditions give preliminary information but real-time data is what counts.
The microbiological side is a separate body of work: microbial enumeration, testing for specified microorganisms, sterility testing for sterile products and the bacterial endotoxin test. For sterile products, packaging integrity and sterilisation validation also come into play.
Data integrity has moved to the centre of inspections in recent years. It is not enough for the result to be correct; the raw data must be traceable, unalterable and auditable. Validation of electronic record systems has therefore become an inseparable part of analysis.
Method groups in this field
Identity and assay
HPLC, UPLC, UV-Vis spectrophotometry, titration, identity confirmation by FTIR.
Impurities and residues
Related impurities, degradation products, residual solvents (GC headspace), nitrosamine screening.
Elemental impurities
Determination of elemental impurities by ICP-MS and ICP-OES and the associated risk assessment.
Physicochemical
Dissolution and disintegration, hardness, friability, uniformity of dosage units, water content (Karl Fischer), particle size.
Microbiological
Microbial enumeration, specified microorganisms, sterility, bacterial endotoxin, preservative efficacy.
Stability
Real-time and accelerated stability studies; photostability and in-use stability.
Commonly confused in this field
- The pharmacopoeial method is binding; an alternative method may be used only by demonstrating its equivalence.
- A result produced without method validation cannot be used in a licence dossier.
- Accelerated stability does not replace real-time data; shelf life is supported by real-time results.
- The dissolution profile is sensitive to apparatus type, agitation rate and medium; the conditions must be complete in the report.
- A shortcoming in data integrity produces a non-conformity in an inspection even when the result is correct.
- A change of formulation or supplier requires analytical comparison and, where necessary, a new stability study.
Which decisions these results support
- The licence dossier and batch release
- Determining shelf life and storage conditions
- Method transfer and method validation
- Incoming raw material acceptance control from suppliers
- Assessing the effect of a formulation change on bioequivalence
Related Tests
Frequently Asked Questions About Dissolution
Does dissolution testing replace bioequivalence?
Generally not. A waiver from a bioavailability study is possible under certain conditions and for certain product classes, but this is a special case assessed against the regulatory authority's guidance.
Why is the medium degassed?
Dissolved gas forms bubbles that stick to the tablet surface and slow dissolution. This creates a spuriously low result and high variability, so degassing is a mandatory step.